Medical Device Recall: A Facility Response Checklist
Medical device recall notice received? Check affected units, interim use, corrective action, staff communication and return-to-use records.
A facility receives a manufacturer email warning that a device model may need a software update, inspection, replacement part, or removal from use. The notice may name serial numbers and a deadline. Staff must decide whether their exact equipment is affected, who can safely use it in the meantime, and how to document the response. The first step is not to panic or ignore the notice. It is to verify the device identity, read the specific instructions, and coordinate a controlled response.
This article is a practical guide to medical device recall and field safety notice response for healthcare facilities. It is distinct from our equipment inventory guide, which explains what information to record before a notice arrives. The inventory is the tool that lets a facility find affected assets. This guide focuses on what to do when an alert actually arrives. It is not legal advice, and the manufacturer's notice plus applicable local requirements take precedence over a general article.
A recall does not always mean the device must be returned
The word “recall” can cover different corrective actions. The World Health Organization's field safety notice guidance explains how manufacturers communicate corrective actions and safety information to device users. A notice may call for inspection, software correction, relabeling, part replacement, revised operating instructions, or taking equipment out of service. It is unsafe to assume every notice means immediate disposal. It is equally unsafe to assume that a device can remain in use because it still powers on.
The action depends on the exact notice, model, serial or lot range, risk, and clinical use. A manufacturer may ask users to stop use immediately, continue with additional checks, or schedule a service visit. Read the full communication rather than a forwarded subject line. If language is unclear, contact the manufacturer or authorized representative and document the clarification.
Step 1: Confirm that the notice is genuine and complete
Identify the sender, manufacturer, date, document version, affected products, and contact route. A notice forwarded by a colleague may omit attachments or later updates. Use the manufacturer's official support channel or an official regulatory alert page to confirm the current instructions. Do not click an unfamiliar link or install a file merely because an email claims to be urgent. Connected equipment also creates cybersecurity considerations.
Keep the original notice in a controlled record. Note when the facility received it and who was assigned to review it. If there are several locations, send the notice to the right clinical, biomedical, procurement, and safety roles. The person who first reads the email may not be qualified to decide whether care can continue. A simple escalation route prevents the alert from being lost in a shared inbox.
Step 2: Match the exact device or lot
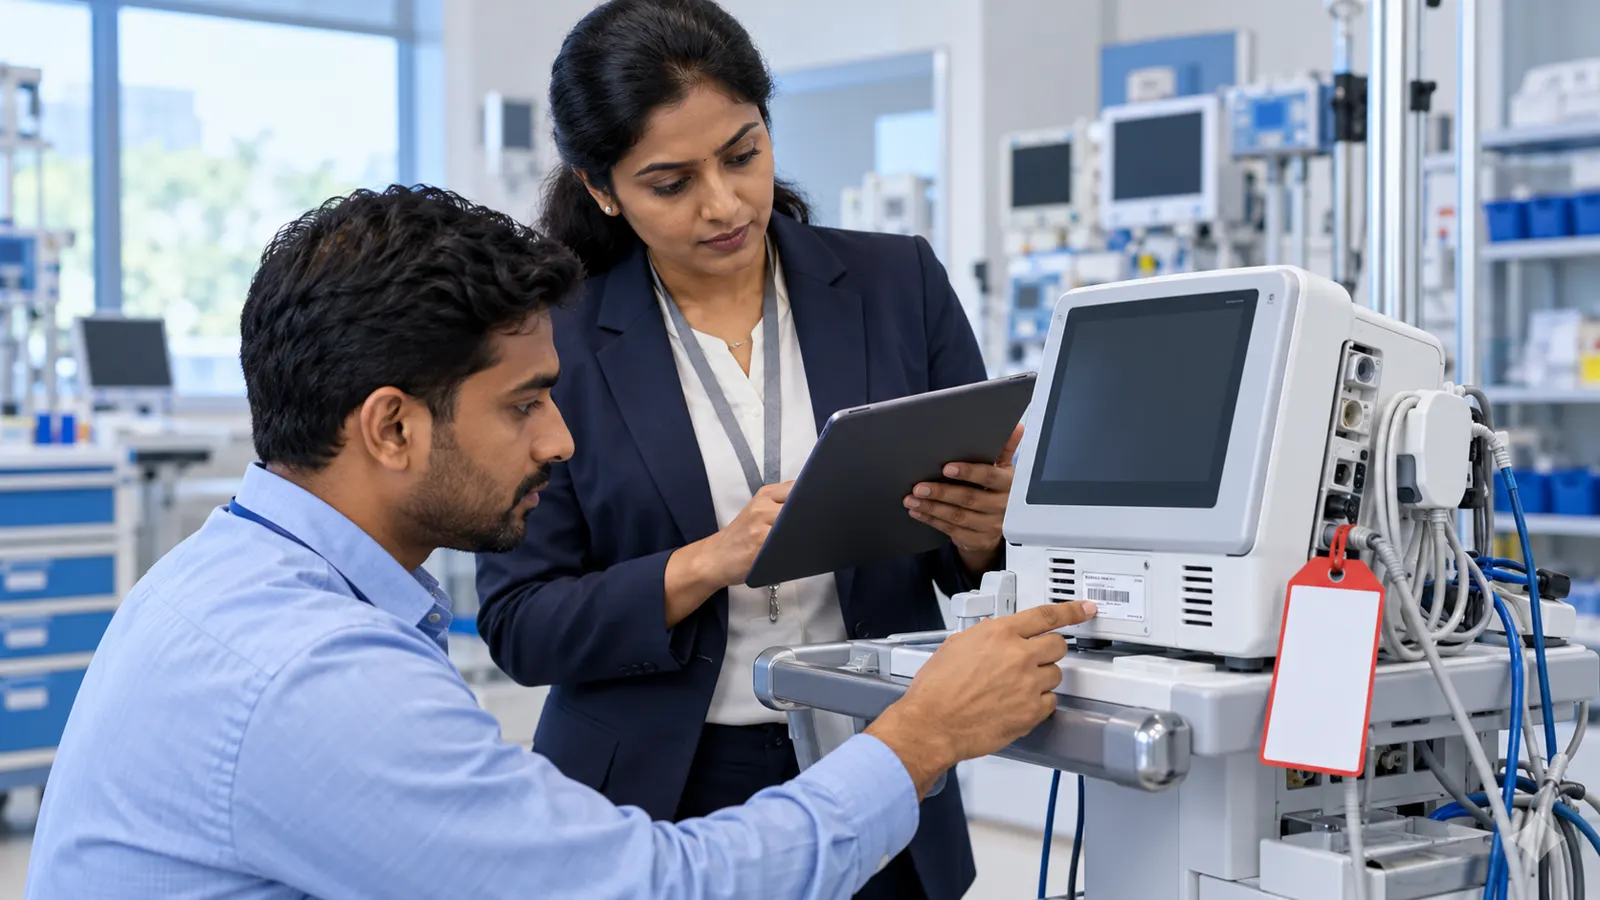
Compare the notice against your asset register: manufacturer, model, serial number, software version, accessory ID, and location. Some notices apply only to a subset of units. Others include accessories or consumables rather than the base machine. Do not rely on a generic name such as “ultrasound machine” or “treadmill.” Physically confirm the label when feasible and record who checked it.
The WHO medical equipment inventory guidance explains why an up-to-date inventory supports alerts and recalls. If the asset list is incomplete, search clinical rooms, storage, mobile equipment pools, and recent service records. A missing item should trigger an investigation, not an assumption that the facility never had it.
Step 3: Decide interim use with qualified people
If the notice says stop use, follow it and mark the device unavailable in the inventory and at the point of use. If the notice permits limited use with conditions, have the relevant clinical and technical leaders assess whether those conditions can be met. Document the decision, who made it, and the temporary instructions for staff. Do not let a general article overrule manufacturer instructions or the facility's safety policy.
Think about patients and workflow. Is there another device or referral pathway? Does the affected machine support urgent care? Can appointments be rescheduled safely? If a device has already been used in a way described by the alert, a clinical team may need to review records or contact patients. That decision requires the notice, clinical context, and appropriate governance. An equipment vendor should not make patient-care decisions alone.
Step 4: Communicate clearly to users
A warning is useful only if the people who operate the device know what changed. Tell them which exact unit is affected, what they may or may not do, and how to report any problem. Use the local asset ID and location. Avoid vague messages that make staff think every similar device is unsafe. Place a clear physical tag or controlled status label where appropriate, and update scheduling or booking systems if the device is unavailable.
If revised instructions are issued, provide training before expecting staff to follow them. Confirm that night and weekend teams receive the same message. Record who was notified and when. Do not rely on one verbal handover. The installation and handover checklist describes how to document operator training when a device first arrives; recall-related instruction changes deserve the same care.
Step 5: Arrange the corrective action
The manufacturer may send an engineer, provide an approved replacement component, issue software, or require a return. Confirm who will perform the work, what qualifications or authorization they need, and what evidence will be supplied afterward. Do not let untrained staff perform a repair or install an unofficial patch to satisfy a deadline. For a connected device, coordinate any software change with the facility's IT and cybersecurity processes.
Keep ticket numbers, the date of action, parts or versions changed, inspection and performance results, and the engineer's report. If the corrective action is delayed, document the interim status and next escalation. Our warranty versus AMC guide explains why service responsibility should be clear before a fault or alert occurs. A recall may involve a manufacturer even when a separate provider handles routine maintenance, so confirm the right route.
Step 6: Verify before returning the device to use
Completion of a service visit is not always the same as safe release to use. Check that the required correction was actually performed on the affected unit, the correct version or part is present, and any mandated performance or safety tests were completed. A qualified person should approve return to use according to the facility's policy and the manufacturer's instructions. Update the asset status and retain the evidence.
If staff were instructed to change a clinical workflow, confirm that the current notice has been distributed and training completed. If the manufacturer later updates the notice, review whether the previous action still satisfies it. A facility needs an audit trail from receipt of alert through closure, including any units that were found unaffected and why.
What should an alert record contain?
Create a short record for each safety notice with:
- Manufacturer and notice title, issue date, and version.
- Date received and source used to verify authenticity.
- Affected model, serial or lot range, and software version.
- Local asset IDs, locations, and responsible departments checked.
- Immediate use decision and the person who approved it.
- Staff and clinical teams notified.
- Service ticket, corrective action, and completion evidence.
- Tests or checks before return to use.
- Patient-impact review if required by the notice and clinical team.
- Closure date, approver, and location of final records.
This is an operational checklist, not a substitute for local reporting duties. Regulatory requirements differ by country and device. In India, the Central Drugs Standard Control Organization (CDSCO) publishes medical-device rules and safety communications. Facilities should follow applicable Indian requirements and the manufacturer's field safety communication, with qualified compliance advice when needed. The CDSCO Medical Devices Rules are a primary reference, but individual obligations depend on the situation.
Do not confuse recall response with routine maintenance
Routine preventive maintenance is planned around device instructions and risk. A recall is an alert about a specified product issue that calls for a particular response. A device may be current on its AMC visits and still be affected by a recall. Conversely, a device that has no active service agreement still needs appropriate action if an alert applies. Use the AMC maintenance checklist for ongoing service processes, and keep recall records linked to the same asset history.
Facilities can improve readiness before any alert arrives by maintaining serial numbers, a named safety-contact role, current vendor contacts, and a clear out-of-service procedure. This is easier than reconstructing ownership and location during an urgent notice. If you need to review the equipment records and service pathway across a facility, contact Regenis with the device categories and operating setting.
Frequently asked questions
Does a recall mean every unit must be removed?
No. The notice may require correction, inspection, updated instructions, or removal for a defined set of devices. Follow the exact product-specific instructions.
Can staff continue using a recalled device until the engineer arrives?
Only if the notice permits it and the facility's qualified clinical and technical team confirms the conditions can be met. Do not infer permission from the device appearing to work.
How do we know whether our equipment is affected?
Match the manufacturer, model, serial or lot numbers, and software version in the notice to the physical device and asset register. Record the result.
Who should close a recall action?
A person or team authorized by the facility should verify the corrective work, required tests, communication, and safe return-to-use decision, then document closure.
Are FDA recall rules the same as Indian requirements?
No. FDA material can explain concepts, but Indian facilities must follow applicable local requirements and the manufacturer's notice. Seek qualified advice for specific reporting duties.